A special process is not audit jargon - it is the factory's most misunderstood risk category
If your process output cannot be verified by inspecting the finished product, you are dealing with a special process - where failure does not show up in the component, but in the patient.
If the output of your process cannot be fully verified through inspection or testing of the finished product, you are dealing with a special process. Here is the uncomfortable truth: in precisely these processes, a failure does not show up in the component - it shows up in the patient.
Sterilisation, cleanrooms and sterile barriers have something in common. You cannot simply look at a product and declare it sterile. You cannot open the packaging to confirm that the barrier has been maintained - opening it destroys it. Compliance in these processes is not measured in the finished product; it is built, validated and maintained over time. That is why clause 7.5.6 of ISO 13485:2016 gives them specific treatment.
What clause 7.5.6 actually requires
The standard is direct: processes whose output cannot be verified through subsequent monitoring or measurement must be validated. In ISO 13485, validation does not mean running the process once and approving it. It means demonstrating, against defined and documented criteria, the ability to produce conforming output consistently and repeatably. The clause requires criteria for review and approval, qualification of equipment and personnel, the use of defined methods and procedures, record requirements and, where applicable, revalidation.
Why verification is not enough
Verification answers, "Does this component conform?" Validation answers, "Can this process consistently produce conforming components?" In a special process, the first question cannot be answered without destroying the product - or there may simply be no observable answer. That leaves the second. Treating a special process as though final inspection were sufficient is the conceptual error that can bring down entire quality systems during an audit.
The link to ISO 14971: risk determines the level of rigour
Not every special process requires the same validation effort. Risk determines the intensity. ISO 14971 provides the logic: the greater the process's contribution to potential patient harm, the greater the rigour required in validation, monitoring and revalidation. This means that a validation strategy does not start with a generic template - it starts with the device's risk analysis. Connecting clause 7.5.6 to ISO 14971 turns validation from a "documentation obligation" into "defensible risk control".
Traceability: the thread that holds everything together during an audit
Identified risk → process specification → testing/qualification → acceptance criterion → ongoing monitoring. When an ANVISA, FDA or European notified body auditor follows this thread, each link must connect to the one before it. The traceability matrix demonstrates that validation was not an isolated event, but a decision based on risk - from beginning to end.
The most common strategic mistake
Validating special processes in silos: sterilisation with one team, the cleanroom with another and packaging with a third - each following its own logic, without communicating. Yet the sterile product that reaches the patient is the result of the combination of these processes. An intact sterile barrier cannot compensate for inadequately validated sterilisation, and the reverse is also true. Special processes are not validated by department. They are validated across the risk chain.
Documentary compliance gets an audit approved. Real process control protects the patient. The two are not synonymous - and the difference tends to emerge at the worst possible moment.
In your organisation, does special process validation start with a template or with the device's risk analysis?
At HSC Global, we help medical device manufacturers structure special process validation as an integrated whole - from risk to operations - with traceability that can be defended in ANVISA, FDA and MDR audits.
Explore the Special Process Validation for Medical Devices course or speak to our team.
Let us discuss your special processes.
#MedicalDevices #ISO13485 #SpecialProcess #ProcessValidation #ISO14971 #RegulatoryAffairs #QualityManagement #MedicalDevices #ANVISA #QMSR #MDR #RDC751 #HSCGlobal
More quality to explore
Each article connects regulatory changes to what they mean for your day-to-day work.
 Quality
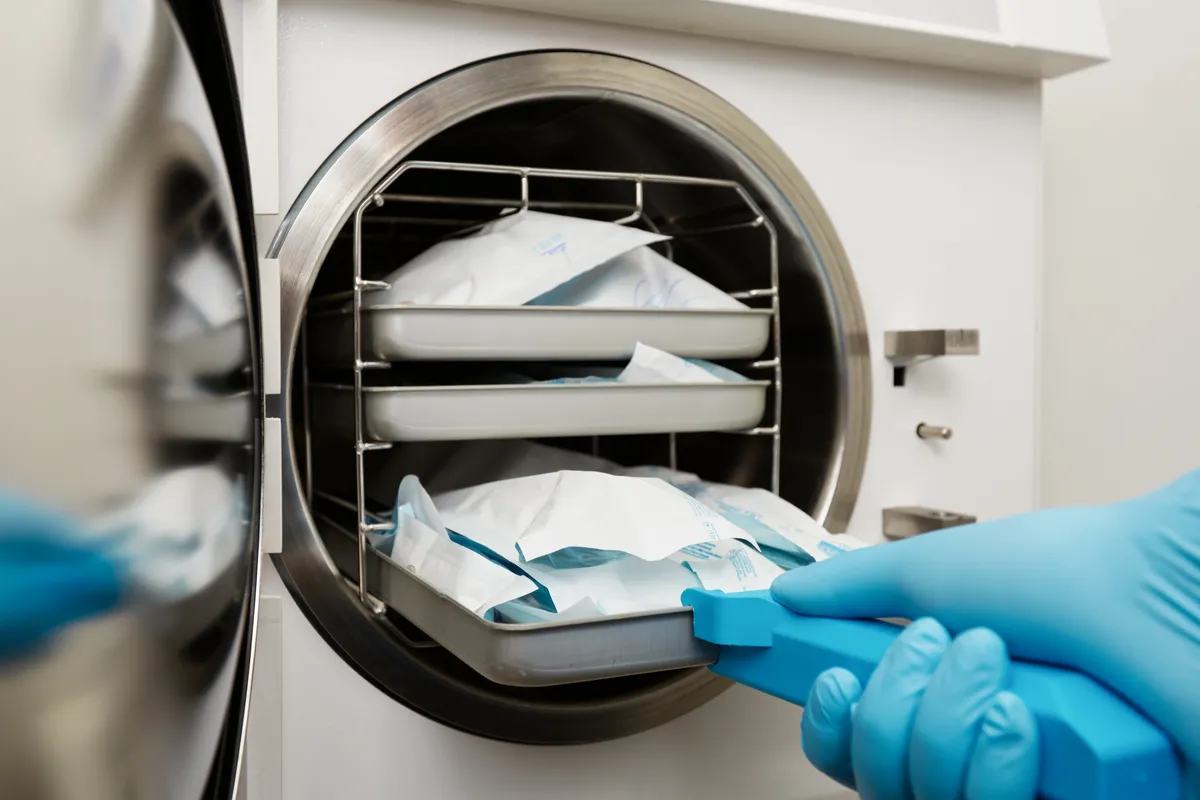
Your sterilisation cycle is validated. But did you validate the right product under the right worst-case conditions?
An approved cycle shows that the equipment delivers the process - not that the most challenging device, in the most demanding load, achieves the required SAL. This confusion is one of the sector's costliest gaps.
 Quality
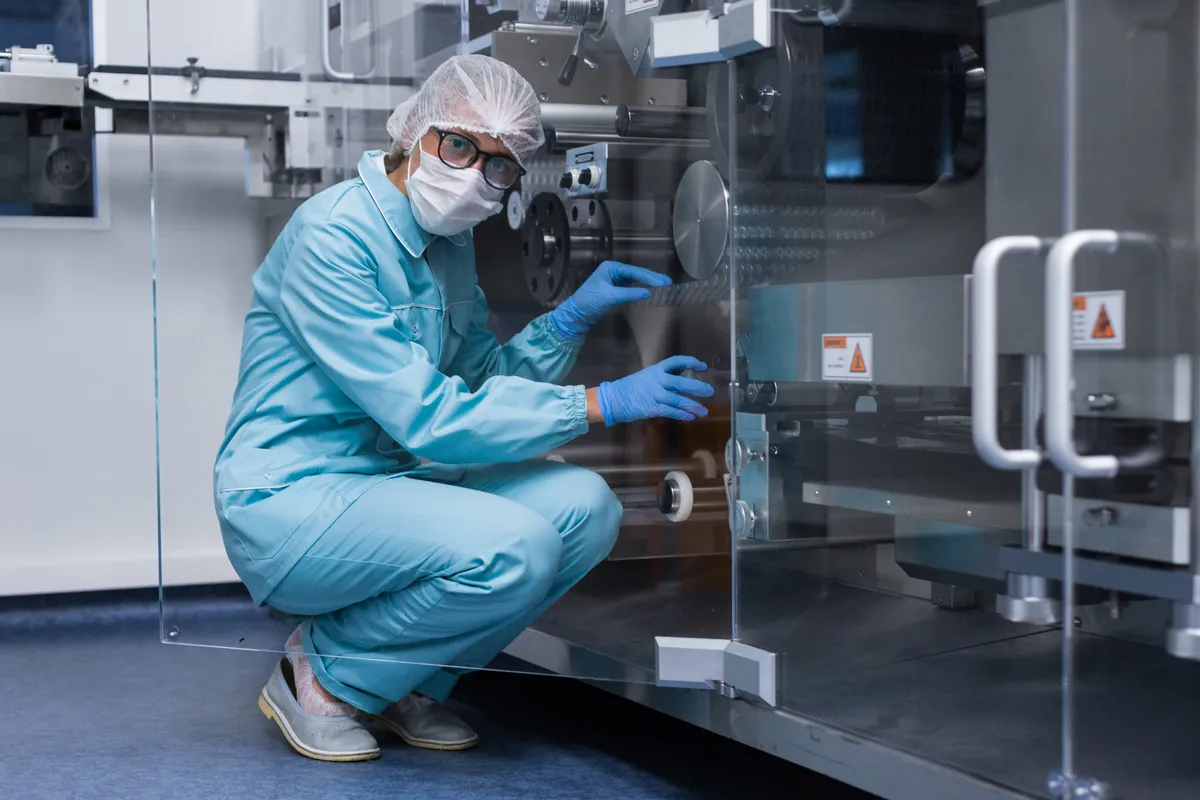
Packaging does more than protect the product. It maintains sterility - and that takes different engineering
Physically protecting a product and maintaining a sterile barrier throughout its shelf life are distinct objectives requiring distinct validations. Sterility only reaches the patient if the packaging preserves it.
 Quality
Four Regulatory Requirements, One Validation System (QMSR, MDR and RDC 751)
ISO 13485, FDA QMSR, MDR 745/2017 and RDC 751/2022: treating each framework as a separate project multiplies costs and weakens your case. The solution is one system interpreted through several regulatory frameworks.
Comments
No comments yet. Be the first to comment.