ANVISA
RDC 591/2021
- Staggered deadlines by risk class
- Data in the ANVISA system
DeadlineThe current timetable for each class is confirmed during the initial assessment.
Unique device identification
HSC structures your portfolio's entire UDI cycle: coding strategy, labelling, data validation and submission to ANVISA, the FDA and the European Union.
Why now?
The staggered deadlines under RDC 591/2021 are already in force by risk class, and the FDA and European Union have required UDI for years. Without compliance, the product cannot circulate.
ANVISA
United States
European Union
The concept
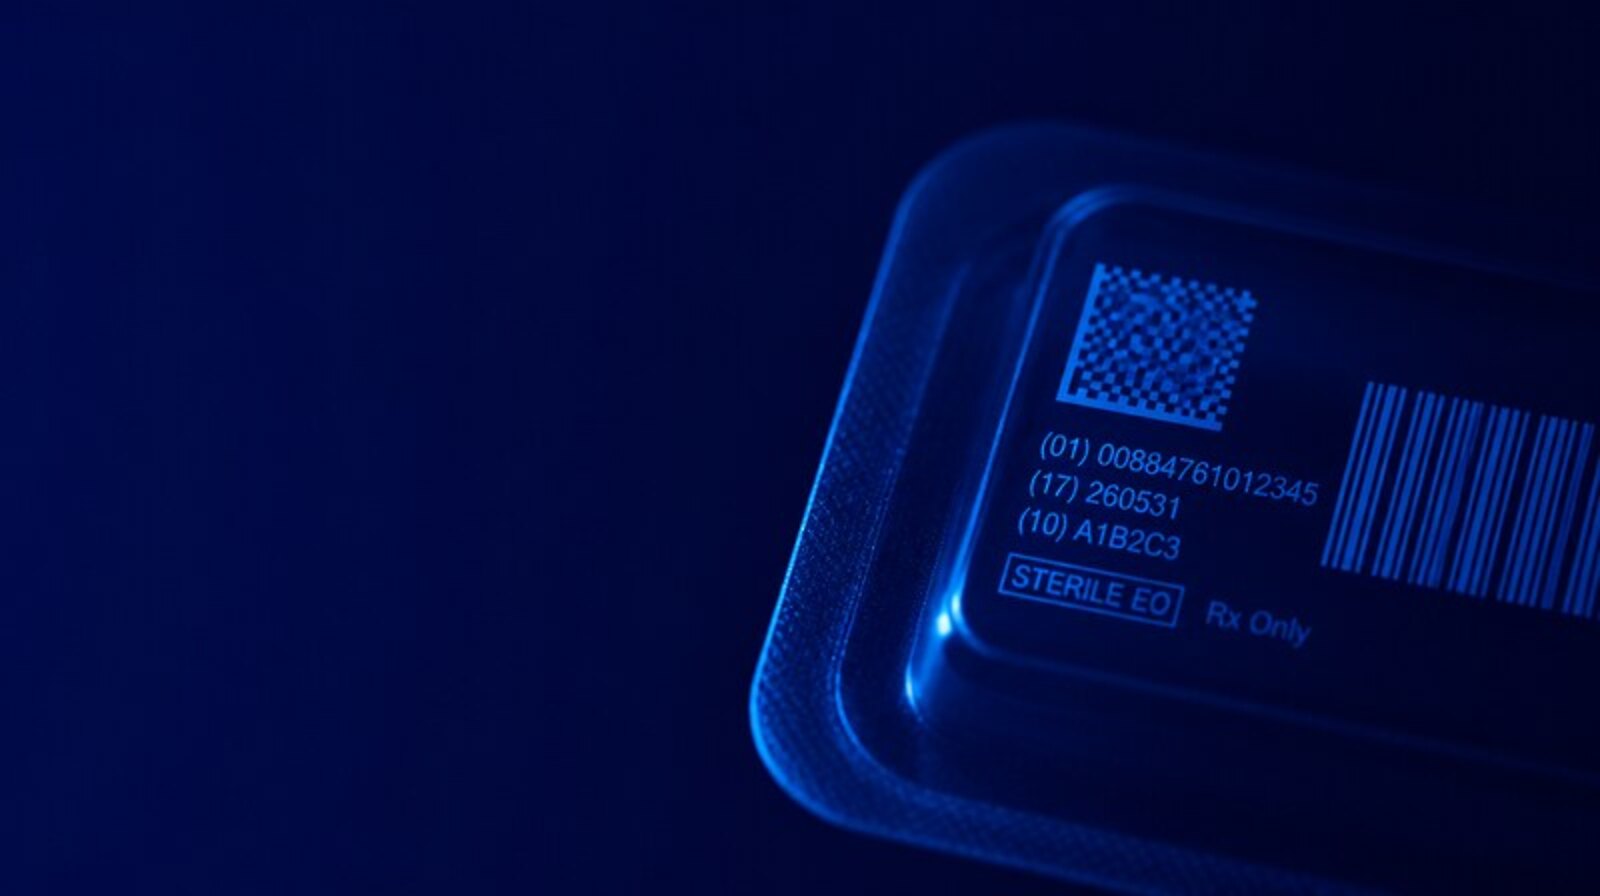
Fixed component associated with the product model and version. It does not change between batches.
Variable component: batch, serial number, expiry date and date of manufacture.
Identifiers are assigned by accredited issuing agencies such as GS1, HIBCC and ICCBBA. Selecting an agency forms part of the coding strategy. For the US market, see our FDA regulatory services.
Service scope
Portfolio assessment, risk classification and mapping of applicable requirements by market and class.
Defining the issuing agency, UDI-DI and UDI-PI architecture by product family, and the approach for kits, configurable devices and software (SaMD).
Marking requirements for the label and the device itself (direct marking where applicable), AIDC and HRI formats, and artwork review.
Preparing the data attributes required by each regulatory database, consistency validation and information governance.
Registration and submission to ANVISA, GUDID (FDA) and EUDAMED (EU), including account and credential management.
UDI procedures incorporated into the quality management system (ISO 13485), change control and ongoing record maintenance.
What you receive
Why HSC?
Recognised by Exemplar Global. Technical expertise extending to UDI implementation.
One partner to implement UDI consistently across all your regulated markets.
Our team already manages labelling, registration and QMS processes. UDI becomes part of your quality system.
We support manufacturers from China, the United States and Europe requiring UDI for ANVISA registration.
Frequently asked questions
Speak to an HSC specialist for an initial assessment of your portfolio's UDI status in the markets where you operate.